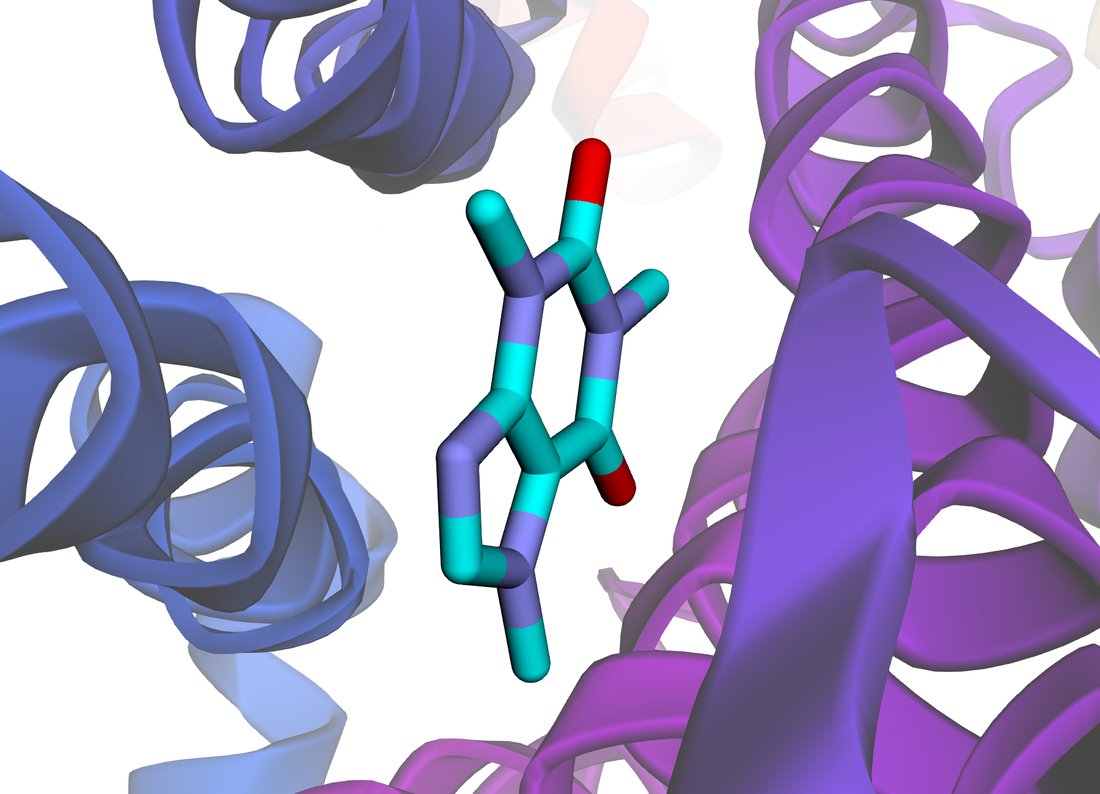
РезультатResultErgebnis
- Molecule
- Caffeine (PubChem CID 2519)
- Target
- Adenosine A2A receptor (PDB 3EML)
- MW / LogP
- 194.19 / −1.03
- TPSA
- 61.82 Ų
- Час розрахунку
- Compute time
- Rechenzeit
- ~9.6s
Як це працюєHow this worksWie das funktioniert
Один і той самий інструмент — будь-яка пара молекула-мішеньOne tool, any molecule-target pairEin Werkzeug — jedes Molekül-Zielprotein-Paar
01
Молекула з PubChem: SMILES, формула, вагаMolecule from PubChem: SMILES, formula, weightMolekül aus PubChem: SMILES, Formel, Gewicht
02
3D-структура білка з RCSB PDB3D protein structure from RCSB PDB3D-Proteinstruktur aus der RCSB PDB
03
AutoDock Vina: реальна фізика зв'язуванняAutoDock Vina: real binding physicsAutoDock Vina: reale Bindungsphysik
04
3D-візуалізація пози в кишені білка3D visualization of the pose in the pocket3D-Visualisierung der Pose in der Bindetasche
У Фазі 2 цей самий пайплайн відпрацював 18 разів поспіль без ручного втручання — на трьох реальних кандидатах проти шести мішеней. Повні дані того прогону — у закритому розділі для інвесторів. In Phase 2, this exact pipeline ran 18 times in a row without manual intervention — on three real candidates against six targets. The full results of that run live in the investor-only section. In Phase 2 lief genau diese Pipeline 18 Mal hintereinander ohne manuellen Eingriff — an drei realen Kandidaten gegen sechs Zielproteine. Die vollständigen Ergebnisse dieses Laufs finden Sie im investorenexklusiven Bereich.
Інструмент докінгуDocking engineDocking-Engine
Розрахунки виконує AutoDock Vina — відкритий (open-source) інструмент молекулярного докінгу, розроблений у Scripps Research (лабораторія Forli). Це один із найпоширеніших докінг-рушіїв у фармацевтичній та академічній індустрії, з тисячами цитувань у наукових публікаціях. Ми запускаємо офіційний бінарник локально, без сторонніх SaaS чи "чорних скриньок" — весь код пайплайна навколо Vina (підготовка молекул через RDKit/meeko, візуалізація через py3Dmol) також можна показати за запитом. The calculations are performed by AutoDock Vina — an open-source molecular docking engine developed at Scripps Research (Forli Lab). It's one of the most widely used docking engines in pharmaceutical and academic drug discovery, with thousands of citations in the scientific literature. We run the official binary locally — no third-party SaaS or black boxes — the surrounding pipeline code (molecule prep via RDKit/meeko, visualization via py3Dmol) can also be shown on request. Die Berechnungen führt AutoDock Vina durch — ein Open-Source-Werkzeug für molekulares Docking, entwickelt bei Scripps Research (Forli Lab). Es ist eine der meistgenutzten Docking-Engines in der pharmazeutischen und akademischen Wirkstoffforschung, mit tausenden Zitierungen in der Fachliteratur. Wir betreiben die offizielle Binärdatei lokal — ohne Drittanbieter-SaaS oder Blackbox-Lösungen — der umgebende Pipeline-Code (Molekülvorbereitung via RDKit/meeko, Visualisierung via py3Dmol) kann auf Anfrage ebenfalls gezeigt werden.
Eberhardt, J., Santos-Martins, D., Tillack, A.F., Forli, S. (2021). AutoDock Vina 1.2.0: New Docking Methods, Expanded Force Field, and Python Bindings. Journal of Chemical Information and Modeling, 61(8), 3891–3898.